Are You Keeping Up With Advances In Medical Bioprinting Technology
September 17, 2019
Coronavirus 2020: Everything Pharmaceutical Personnel Should Know

3D printing technology has advanced rapidly in the last decade. The method has quickly developed into one of the leading solutions for a variety of medical issues. 3D printing provides a cost-effective and efficient way to tailor medical treatment to individual patients.


Recently, the FDA has accepted many 3D printing methods for medical and pharmaceutical applications. As a new technology, 3D printing is quickly improving in every field. New innovations are being submitted to the FDA for approval every day. 


Medical Applications of Medical Bioprinting

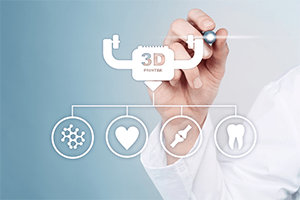
There is a wide range of applications for medical bioprinting technology. Everything from artificial bone transplants to pharmaceutical drugs is manufacturable on a 3D printer. 


Here are a few use cases highlighting why many companies are looking at the medical applications of 3D printing:

  • Quickly manufacture replacement knees
  • Bioprinting tissues using specialized technology to layer living cells (known as bio-ink)
  • Drugs manufactured especially for fast absorption by the body


As a new field, the applications for bioprinting are continually growing. In the future, 3D printing methods are expected to be used to print skin cells directly onto a burn patient’s wounds. 


The idea of printing fully-functional organs isn’t that far off either. Researchers are already working on methods to make this possible. 

But as this technology matures, so too are the regulations that oversee it.


Why It’s Important to Keep Your Bioprinting Methods Up To Date and Compliant

With new medical discoveries made every day, it’s essential to use the most current medical bioprinting methods. All new technologies carry some level of risk. The FDA audits new medical treatments, but it cannot guarantee that they are safe. 


Keeping up to date with the latest practices in 3D bioprinting allows you to make use of the safest and most effective pharmaceutical methods. Staying ahead of the curve helps keep your business competitive and your patients or customers safe. 


Keep Your Existing Processes Current and FDA-Compliant

The Center for Professional Innovation and Education (CfPIE) is your ultimate resource for staying up to date on the latest changes in the pharmaceutical, biotech, medical device, skincare, and cosmetics. 


At CfPIE, we provide training courses that work to help you build FDA-compliant processes that will increase organizational accountability, documentation practices, and other health and safety mandates.


Check out some of our pharmaceutical and biotech and medical device training courses to ensure that your knowledge is modern and relevant.


Do you have questions about our pharma health and training courses? 
Contact us today to learn more.


paper_plane

Need more information on our life-science training course?

Get In Touch With Us Today!

Contact Us

Blog Categories

October 27, 2026
Our analysis identified quality leadership and culture as a foundational systemic gap. When senior management does not give the quality unit sufficient authority, resources, and organizational support, deficiencies rarely stay confined to one procedure or department. They can affect investigations, facility maintenance, production decisions, corrective and preventive action (CAPA) effectiveness, and timely risk escalation throughout the pharmaceutical quality system (PQS). What the Warning Letters Revealed The Warning Letters reviewed for this analysis repeatedly showed three related patterns: ineffective quality-unit oversight, inadequate management attention to known deficiencies, and corrective actions that addressed individual observations without resolving their underlying systemic causes. 1. Ineffective Quality Unit Authority Quality units (QUs) commonly exist on paper but often lack the authority, resources, or competence to enforce current good manufacturing practice (cGMP) compliance. These limitations lead to a host of issues, from failing to investigate discrepancies to allowing unsafe production practices to continue. For example, in a June 2026 Warning Letter , the FDA found that Excelvision, a Fareva facility, was disinfecting critical equipment components with sporicide rather than sterilizing them between batches. The pharmaceutical manufacturer responded by stating that some components cannot be sterilized due to machine design. The company also claimed they would use sporicide between each batch, increase microbiological sampling, and swab equipment, but the FDA deemed these actions unacceptable substitutes for sterilization. FDA concluded that Excelvision's QU either lacked the proper authority to carry out its responsibilities or had insufficiently implemented them. The company must develop a remediation plan to ensure its QU has the authority and resources to function effectively, among other requirements. 2. Inadequate Executive Oversight and Resourcing Warning Letters commonly point to senior leaders failing to resolve deficiencies and systemic flaws, rather than exercising the oversight that cGMP compliance requires. Without that oversight, employees may find workarounds or hide issues that executive leadership failed to address proactively. Some leaders do not give QU the funds to ensure maintenance programs are followed, compromising product safety and ultimately forcing large-scale remediation or heightened scrutiny. Consider the June 2026 Warning Letter issued to Huons Co., Ltd . FDA found that a microbiology team leader directed staff to discard bioburden plates that showed contamination, and changed timestamps to create backdated documentation. The team leader also admitted to cutting pages out of a logbook and replacing them with fabricated ones, instructing an analyst to backdate and omit bioburden testing results. Huons must conduct a comprehensive investigation into the scope of the falsification, perform a risk assessment of the quality of the affected drugs, and commit to two years of third-party audits to verify its corrective actions, among other requirements, to address the violations. In the Warning Letter to Excelvision, the FDA found that the company had built parts of its sterile manufacturing areas, including filling rooms and compounding rooms, with ceiling and partition materials that could not be waterproofed or decontaminated. The facility experienced repeated water intrusion, and mold was detected in cleanrooms. Now the company must implement a CAPA plan ensuring vigilant oversight of equipment and facility conditions, including making necessary repairs and upgrades. 3. Reactive CAPAs and Repeated Deficiencies Throughout our research, we concluded that leadership may view recurring problems as isolated incidents that can be fixed by superficial corrective and preventive action (CAPA). Companies commonly retrain employees rather than finding and fixing the root cause. For example, in a December 2025 Warning Letter to Apothecary Pharma, LLC, the FDA found that retraining on aseptic technique failed to address specific behaviors that had caused the original contamination risk. Putting a temporary “band-aid” over problems may keep production moving, but it won't prevent recurring deficiencies. A superficial approach to problem-solving can also go along with reactive compliance, where companies act only after regulators intervene. In both the Excelvision and Huons Warning Letters, for instance, the FDA recommended a voluntary recall before either company committed to one, rather than deciding to recall on their own as a proactive measure.  Why This Is a Systemic PQS Weakness
5 Systemic Training Gaps That FDA Warning Letters Keep Revealing
October 20, 2026
FDA warning letters are often the result of inadequate or generic training. Learn more about five FDA systemic training gaps and how to fill them.